AMG 193 PRMT5 Inhibitor Targets MTAP-Deleted Solid Tumors via Synthetic Lethality
Copy code to add this image to your page
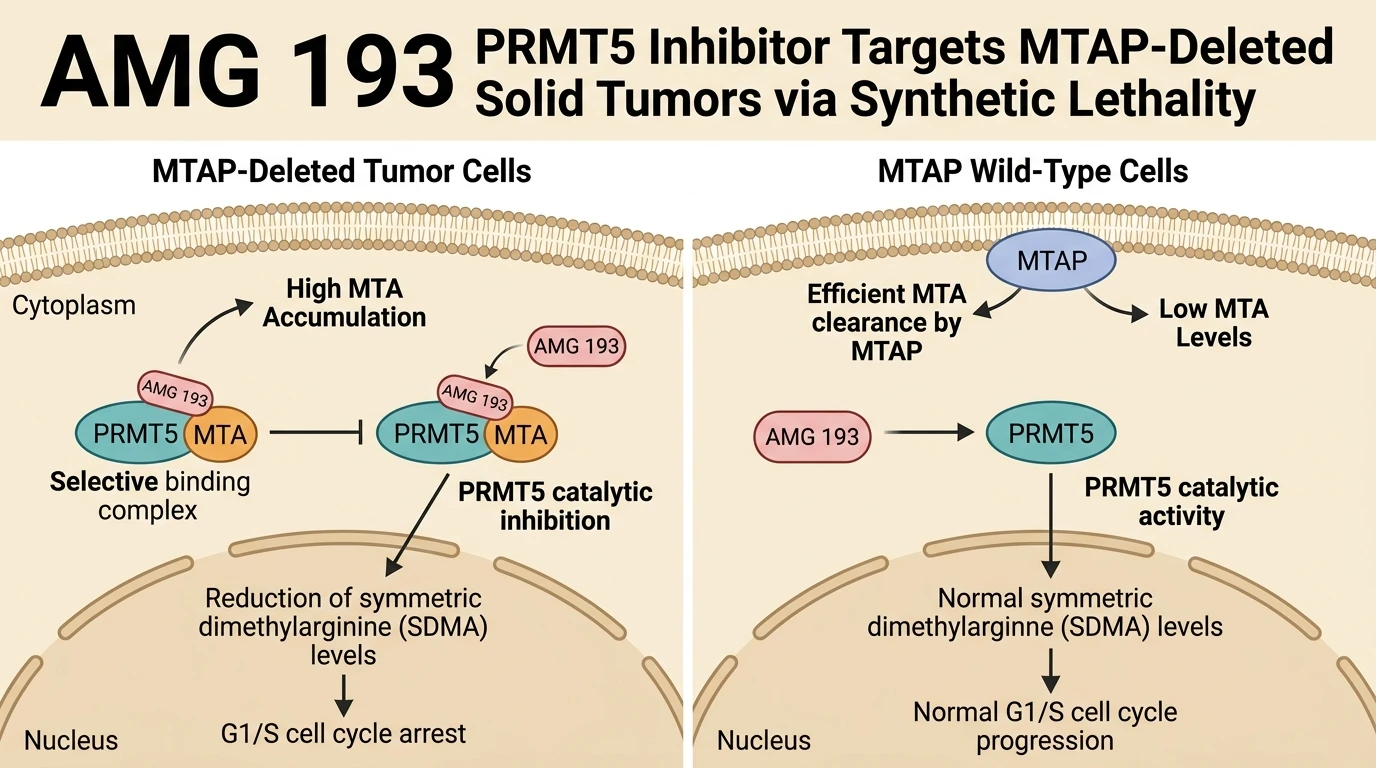
Deletions of the methylthioadenosine phosphorylase (MTAP) gene occur in approximately 10% to 15% of human malignancies, frequently co-deleted with the adjacent tumor suppressor CDKN2A on chromosome 9p21. MTAP plays a key metabolic role in the adenine and methionine salvage pathways by catalyzing the cleavage of S-methyl-5'-thioadenosine (MTA) into adenine and 5-methylthioribose-1-phosphate. Homozygous deletion of MTAP abolishes this enzymatic activity, leading to marked intracellular accumulation of MTA. This metabolic byproduct shares structural homology with S-adenosylmethionine (SAM), the primary methyl donor utilized by protein methyltransferases. Excess intracellular MTA binds to the catalytic site of protein arginine methyltransferase 5 (PRMT5), a type II arginine methyltransferase responsible for symmetric dimethylation of arginine residues on histone tails (e.g., H3R2, H4R3) and non-histone substrates, including ribonucleoproteins involved in pre-mRNA splicing (such as SmD1, SmD3, and RBM15).
Although MTA binding partial inhibits PRMT5 catalytic efficiency, it creates a unique structural alteration that exposes an allosteric pocket specific to the PRMT5–MTA complex. This biochemical dynamic establishes a classical synthetic lethal vulnerability exclusive to MTAP-deleted cancer cells. Normal MTAP-wild-type (MTAP-WT) cells maintain low endogenous MTA levels due to rapid enzymatic clearance, leaving PRMT5 predominantly uncomplexed or bound to SAM. Consequently, standard PRMT5 inhibitors that compete non-selectively with SAM disrupt PRMT5 activity indiscriminately in both malignant and healthy tissues, precipitating dose-limiting toxicities, notably severe bone marrow suppression and grade ≥3 anemia.
AMG 193 was engineered specifically as an oral, MTA-cooperative PRMT5 inhibitor designed to exploit this metabolic discrepancy. Rather than competing directly with SAM or attempting to bind native PRMT5, AMG 193 selectively targets and stabilizes the pre-formed PRMT5–MTA complex. By occupying an adjacent binding site created upon MTA engagement, the AMG 193 PRMT5 Inhibitor locks PRMT5 in an inactive conformation. This cooperative binding mechanism yields orders-of-magnitude higher affinity for PRMT5 in MTAP-deleted cellular contexts compared to MTAP-WT cells, establishing a wide therapeutic window.
Inhibition of the PRMT5–MTA complex by AMG 193 rapidly blocks methyl group transfer to key target substrates, inducing a dose-dependent reduction in global symmetric dimethylarginine (SDMA) levels. Attenuation of SDMA modification disrupts RNA splicing fidelity, triggering aberrant intron retention and exon skipping in essential transcripts such as MDM4. This splicing dysfunction leads to p53 pathway activation in p53-wild-type systems, induction of DNA damage markers (e.g., γ-H2AX), G1/S phase cell cycle arrest, and eventual apoptotic cell death specifically within MTAP-deficient cells.
Preclinical evaluation across diverse cell line panels and patient-derived xenograft (PDX) models confirmed the high selectivity of AMG 193. In MTAP-deleted non-small cell lung cancer (NSCLC), pancreatic ductal adenocarcinoma, esophageal squamous cell carcinoma, and cutaneous melanoma xenografts, daily oral administration of AMG 193 resulted in robust, dose-dependent tumor regressions. Importantly, pharmacodynamic monitoring revealed durable suppression of intratumoral SDMA levels without inducing critical reductions in peripheral red blood cell or platelet counts, validating the targeted hypothesis.
Translational findings translated into early clinical evaluation in a Phase I/Ib trial (NCT05094336) investigating AMG 193 in patients with advanced MTAP-deleted solid tumors. Preliminary safety analysis confirmed a manageable adverse event profile, with low rates of treatment-related grade ≥3 anemia, distinctly contrasting the toxicity profiles observed with non-selective PRMT5 inhibitors. Confirmed partial responses and durable stable disease were documented across multiple tumor types, including NSCLC, pancreatic carcinoma, cholangiocarcinoma, and melanoma, demonstrating proof-of-mechanism for MTA-cooperative targeting in human clinical settings.
Therapeutic Evaluation and Pharmacological Characterization
Copy code to add this image to your page
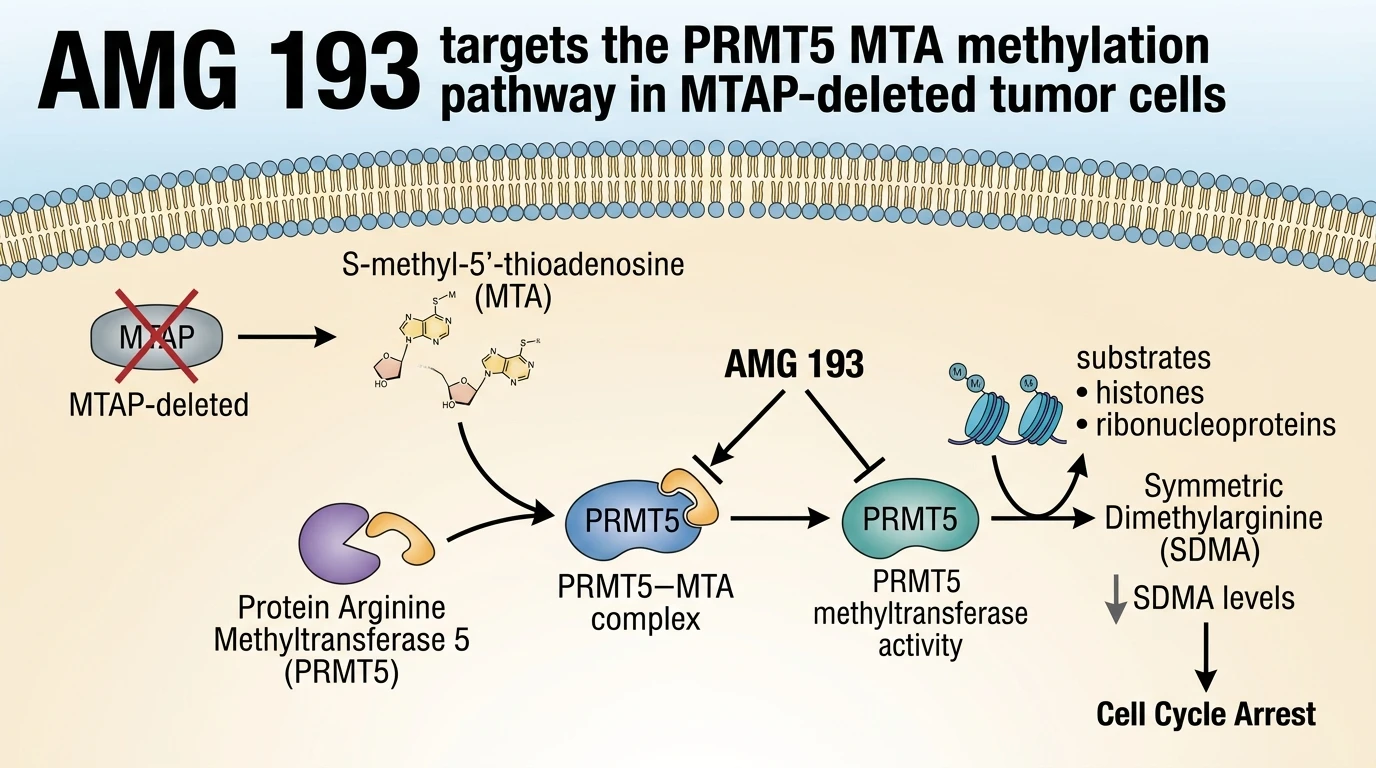
AMG 193 is an oral, MTA-cooperative PRMT5 inhibitor targeting MTAP-deleted cancers through synthetic lethality. In MTAP-deficient cells, accumulated methylthioadenosine (MTA) forms a PRMT5–MTA complex. AMG 193 selectively binds and stabilizes this complex, blocking methyltransferase activity and reducing symmetric dimethylarginine (SDMA) levels. This inhibition triggers RNA splicing defects, cell cycle arrest, and apoptosis in MTAP-deleted tumors while sparing MTAP wild-type tissues. Preclinical and Phase I clinical data demonstrate potent antitumor efficacy and reduced hematologic toxicity compared to non-selective PRMT5 inhibitors.
Selective Binding Traps PRMT5-MTA Complex in MTAP-Deficient Cells
Homozygous deletion of the MTAP gene occurs frequently in human malignancies. This genetic loss disrupts the purine salvage pathway, inducing pronounced intracellular accumulation of S-methyl-5'-thioadenosine (MTA). Accumulated MTA preferentially binds the catalytic domain of PRMT5. The small molecule AMG 193 exploits this metabolic vulnerability by selectively binding the pre-formed PRMT5–MTA complex rather than uncomplexed PRMT5 enzyme. Target residence time increases substantially in this state. Consequently, methyltransferase catalytic activity drops markedly, suppressing symmetric dimethylarginine (SDMA) modification on spliceosomal proteins like SmD3 within tested cellular models.
Dose-Dependent Tumor Regression and Mitigated Myelosuppression In Vivo
Preclinical models demonstrated robust, selective antitumor responses. In murine xenograft studies encompassing MTAP-deleted non-small cell lung cancer and pancreatic ductal adenocarcinoma, the AMG 193 PRMT5 Inhibitor produced dose-dependent tumor regressions. Hematologic toxicity remained notably absent at therapeutic doses. First-generation unselective PRMT5 inhibitors frequently produced severe target-mediated myelosuppression and grade ≥3 anemia due to non-discriminatory inhibition in bone marrow progenitor cells. The MTA-cooperative mechanism preserves normal hematopoiesis.
Clinical Antitumor Activity Across Advanced MTAP-Deleted Solid Tumors
Clinical translation confirmed the differential safety margin. Phase I/Ib evaluation (NCT05094336) in patients with advanced MTAP-deleted solid tumors showed favorable tolerability alongside measurable pharmacodynamic suppression of SDMA. Grade 3 anemia rates remained remarkably low. Confirmed partial responses occurred across diverse tumor histologies, including cholangiocarcinoma, cutaneous melanoma, pancreatic cancer, and non-small cell lung cancer. AMG 193 continuous clinical evaluation remains ongoing.