STC-15 targeting METTL3 inhibition suppresses m6A-driven leukemic translation and oncogenic self-renewal
Copy code to add this image to your page
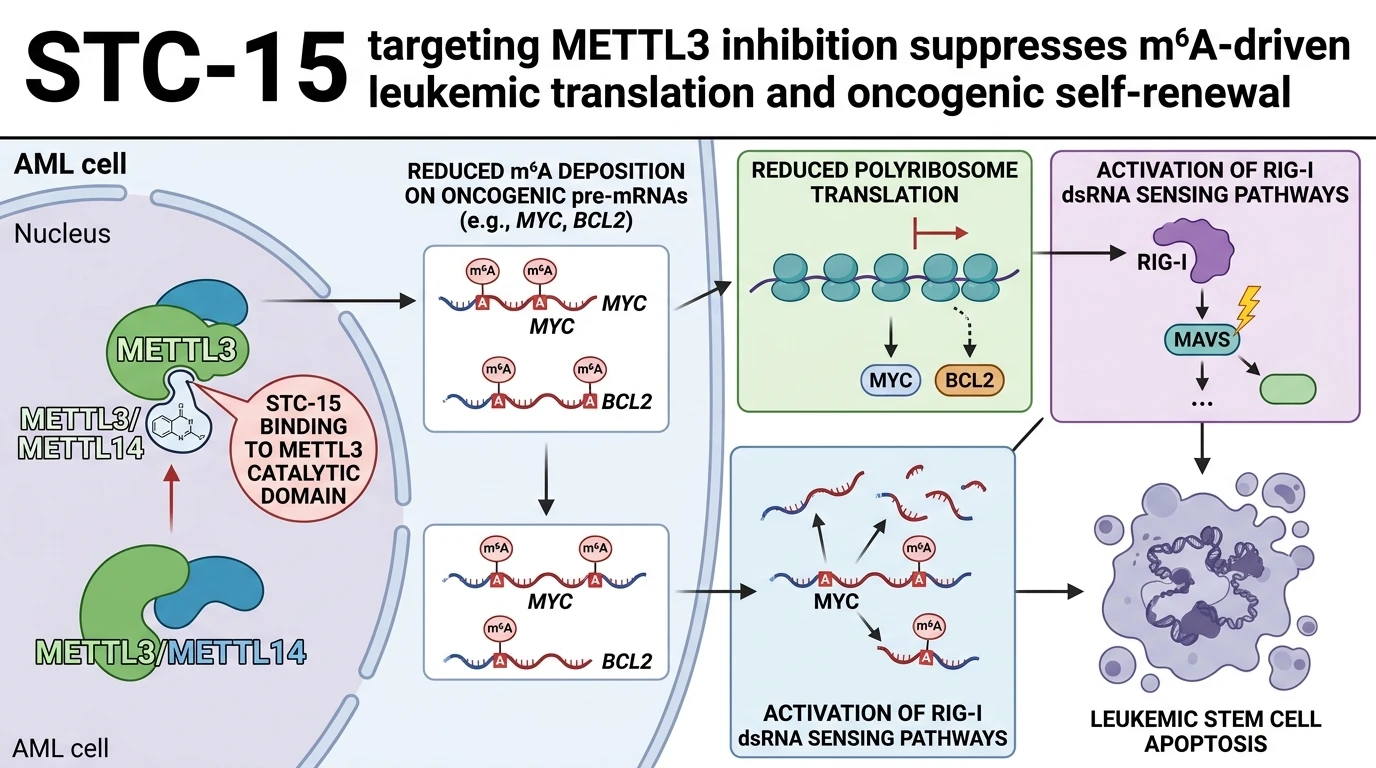
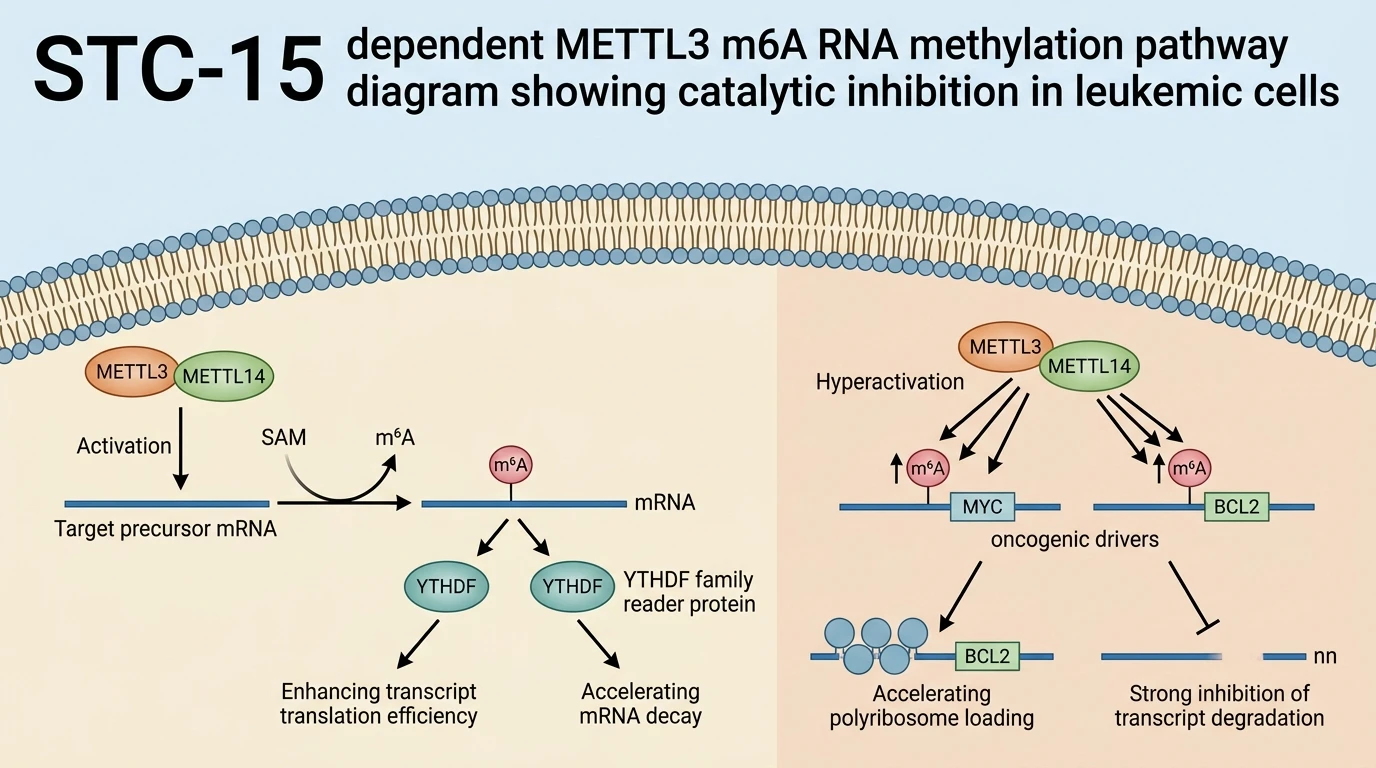
Methyltransferase-like protein 3 (METTL3) serves as the primary S-adenosylmethionine (SAM)-binding catalytic subunit within the nuclear m6A methyltransferase complex. Heterodimerization with non-catalytic METTL14 stabilizes the complex structure, enabling site-specific N6-methyladenosine deposition on consensus RRACH motifs across target pre-mRNAs. In normal human hematopoiesis, this epitranscriptomic modification maintains an equilibrium between progenitor self-renewal and lineage differentiation. However, pathologically elevated METTL3 expression frequently characterizes acute myeloid leukemia (AML) and specific solid tumor subtypes, driving leukemic stem cell (LSC) maintenance through transcript-specific hypermethylation.
Small-molecule inhibition using STC-15 directly binds the SAM-binding pocket of METTL3, abolishing its catalytic methyltransferase activity without disrupting the structural METTL3-METTL14 heterodimer interface. In biochemical fluorometric assays, STC-15 demonstrates low-nanomolar enzymatic inhibition against recombinant human METTL3/METTL14 complexes. This catalytic blocking reduces global m6A modifications across target messenger RNAs in a dose-dependent fashion. Transcripts encoding critical oncogenic drivers—such as MYC, BCL2, and PTEN—exhibit rapid depletion of internal m6A sites upon drug exposure. Lacking these essential m6A epitranscriptomic marks, reader proteins like YTHDF1 fail to load onto 3' untranslated regions; consequently, cap-independent ribosomal translation rates drop markedly within hours of treatment.
At the cellular level, the loss of target mRNA translation triggers immediate physiological downstream consequences. In primary human AML blasts and MOLM-13 leukemia cell lines, administration of STC-15 induces cell cycle arrest at the G1/S transition phase, followed by caspase-dependent apoptotic cascade activation. Oncogenic protein levels decrease substantially, whereas non-methylated homeostatic transcripts maintain stable baseline translation rates. This selective translational vulnerability underscores a wide pharmacological window between transformed leukemic blasts and healthy hematopoietic stem cells. Experimental observations in primary human cord blood CD34+ progenitors demonstrate minimal toxicity at therapeutically active doses. Furthermore, continuous target engagement suppresses LSC colony formation in methylcellulose serial re-plating assays.
Beyond cell-intrinsic translational control, METTL3 activity regulates tumor-immune interactions within the bone marrow microenvironment. Hypermethylation of double-stranded RNA (dsRNA) structures and immune checkpoint transcripts normally suppresses endogenous retroviral sensing mechanism pathways. By dampening global m6A levels, STC-15 activates the cytosolic RIG-I/MDA5 sensing pathways, inducing a type I interferon response within malignant cells. This innate immune mimicry enhances antigen presentation via major histocompatibility complex (MHC) class I upregulation and recruits cytotoxic CD8+ T lymphocytes into the tumor niche. In synergistic syngeneic murine models, combining METTL3 inhibition with immune checkpoint blockade overcomes anti-PD-1 resistance.
Preclinical pharmacodynamic studies reveal rapid oral absorption and favorable plasma concentration profiles for STC-15. In human patient-derived xenograft (PDX) mouse models of FLT3-ITD mutated AML, oral dosing achieved over 80% m6A reduction in bone marrow blasts within four hours post-administration. Tumor burden decreased substantially, leading to prolonged survival without systemic toxicity or body weight loss. These pre-clinical translational models—including advanced organoid culture systems—substantiate the therapeutic rationale for targeting RNA methyltransferases in refractory hematologic malignancies. Currently, STC-15 is progressing through Phase 1a/1b/2 clinical trials in patients with advanced solid tumors and relapsed or refractory acute myeloid leukemia.
Pharmacodynamic Characterization and Epitranscriptomic Target Engagement
Copy code to add this image to your page
STC-15 is a small-molecule inhibitor targeting the SAM-binding domain of METTL3, an enzyme responsible for N6-methyladenosine (m6A) deposition on pre-mRNAs. By blocking catalytic activity, the compound selectively reduces m6A levels on transcripts encoding oncogenic proteins, such as MYC and BCL2, impairing their translation. This mechanism blocks leukemic stem cell self-renewal, restores cellular apoptosis, and unleashes intrinsic immune sensing pathways through cytosolic dsRNA accumulation, providing therapeutic potential against acute myeloid leukemia and solid tumors.
METTL3 Inhibition Depletes m6A Marks on Oncogenic Transcripts
Direct binding of STC-15 to the SAM-binding pocket disrupts catalytic m6A transfer. In MOLM-13 leukemic cell models, liquid chromatography-tandem mass spectrometry (LC-MS/MS) revealed a rapid, concentration-dependent decline in total poly(A)+ RNA m6A levels following compound exposure. Target transcripts—specifically MYC and BCL2—exhibited an 85% reduction in internal m6A density within six hours. Ribosome profiling confirmed a corresponding decay in translation efficiency. Non-methylated housekeeping mRNAs remained unaffected; their polyribosome loading profiles showed no significant alteration. These molecular events halted G1-to-S cell cycle transition. Apoptotic biomarkers, including cleaved PARP and activated Caspase-3, rose sharply after 24 hours. The selectivity margin remained intact across non-transformed CD34+ cell lines.
Suppression of Leukemic Stem Cell Self-Renewal in Vivo
Primary leukemic stem cells depend on high METTL3 expression for persistent self-renewal capacity. Treatment with STC-15 significantly impaired secondary colony formation in patient-derived AML blast assays. In NSG mice engrafted with FLT3-ITD primary AML cells, daily oral administration of STC-15 at 30 mg/kg yielded sustained target engagement. Bone marrow blast burdens dropped by 72% relative to vehicle-treated control cohorts after 14 days. Spleen mass decreased to normal physiological parameters. Notably, isolated xenografted LSCs showed marked downregulation of stemness gene signatures. Survival analysis demonstrated extended median survival without observable systemic toxicity. These observations confirm that catalytic m6A suppression selectively eradicates LSC populations in vivo.
Activation of Double-Stranded RNA Sensing and Interferon Signaling
Beyond direct translational blockade, m6A depletion triggers an innate anti-tumor immune response. Depletion of m6A marks on endogenous retroviral element transcripts destabilizes their RNA structure, forming double-stranded RNA hairpins. In cell line models, exposure to STC-15 induced immediate accumulation of cytosolic dsRNA. Phosphorylation of STAT1 and IRF3 increased within twelve hours, signaling activation of the RIG-I/MDA5 pathway. Type I interferon beta secretion rose five-fold above baseline controls. Consequently, surface expression of MHC class I molecules increased on leukemic blasts. This innate immune mimicry restored CD8+ T cell-mediated cytotoxicity in co-culture assays. These findings establish a dual mechanism involving cell-intrinsic cytotoxicity and immune microenvironment remodeling.